
Preparation method of topiroxostat
A technology of topinostat and solvent, which is applied in the field of drug synthesis, can solve the problems of difficult purification of topinostat crystals, low yield of cyano group substitution reaction, high production cost, etc., and achieve low production cost, cheap price, and three wastes little effect
Patent Information
- Authority / Receiving Office
- CN · China
- Patent Type
- Applications(China)
- Current Assignee / Owner
- Publication Date
- 2021-07-27
Smart Images
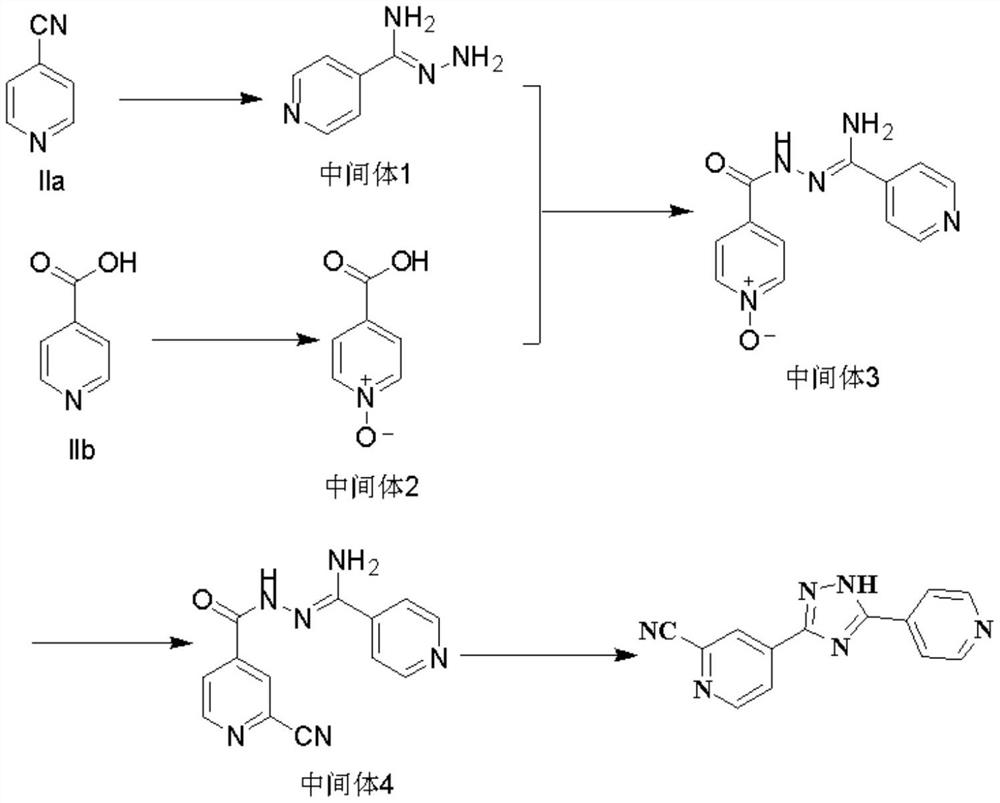
Figure 1 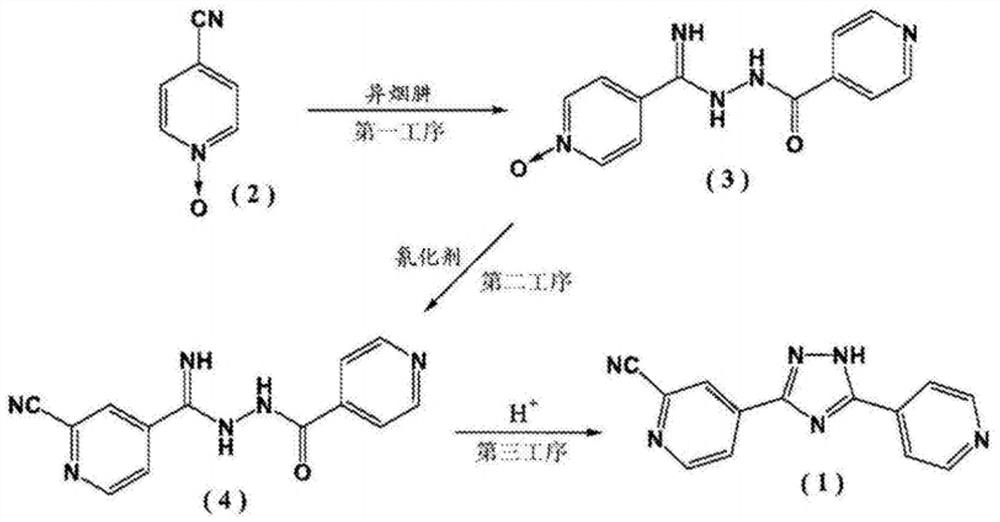
Figure 2 
Figure 3
Abstract
Description
technical field
[0001] The invention relates to the technical field of drug synthesis, in particular to a preparation method of topinostat. Background technique
[0002] Topiroxostat (Topiroxostat, chemical name: 4-[5-(4-pyridyl)-1H-1,2,4-triazol-3-yl]pyridine-2-cyano), manufactured by Japan Fuji Pharmaceutical Co., Ltd. Researched and developed by the company, it was approved by the Japanese Ministry of Health and Welfare in August 2013 and was first launched in Japan. It is a new drug for the treatment of hyperuricemia with gout symptoms. Topinastat is a new type of xanthine oxidase inhibitor, which mainly inhibits the formation of uric acid by linking to the intermediate of oxidative hydrogenation and interacting with amino acid residues.
[0003] The patent CN104411686B applied by Japan Co., Ltd. Fuji Pharmaceuticals in China discloses a preparation method of topinostat:
[0004]
[0005] 4-cyanopyridine N-oxide (2) reacts with isoniazid in the presence of an alkali...
Examples
Embodiment 1
[0034] The present embodiment provides a kind of preparation method of topicastat, and chemical reaction formula is as follows:
[0035]
[0036] The preparation method of the present embodiment topicastat, comprises the following steps:
[0037] a. 4-cyanopyridine is reacted with 80% hydrazine hydrate under heat preservation and stirring for 2.5-4 hours in a solvent and an alkaline reagent, and the intermediate 1 is obtained after post-processing; wherein, the solvent is preferably ethanol, the alkaline reagent is sodium methoxide, and the holding temperature is 25±5°C, the post-treatment is to lower the temperature to 0-10°C, slowly add methyl tert-butyl ether dropwise, control the temperature at 5±5°C, stir and crystallize for 11-13 hours to obtain a yellow solid, filter, and use a small amount of Rinse with methyl tert-butyl ether, then vacuum dry at 25-35°C;
[0038] b. 4-Pyridinecarboxylic acid was heated and stirred under acidic reagent and 30% hydrogen peroxide for...
Embodiment 2
[0043] This embodiment provides a preparation method of intermediate 1, the chemical reaction formula is as follows:
[0044]
[0045] The specific operation process is as follows: Add 833 g (8 mol) of IIa4-cyanopyridine into a 50 L three-necked flask at room temperature, add 4.0 L of ethanol and 17 g (0.31 mol) of sodium methoxide, and stir at 25 ± 5 ° C for 3 h, thin layer Chromatographic detection showed that the raw materials were basically reacted completely; take another 10L four-neck flask, add 4.0L absolute ethanol and 303.6g (6.06mol) of 80% hydrazine hydrate, control the temperature at 25±5°C, keep stirring for 1-2h, thin The transition state reaction was detected by layer chromatography; the temperature was lowered to 0-10°C, and 24L methyl tert-butyl ether was slowly added dropwise, the temperature was controlled at 5±5°C, and the yellow solid was obtained by stirring and crystallizing for 12 hours, which was filtered, and the filter cake was washed with a small ...
Embodiment 3
[0048] This embodiment provides a kind of preparation method of intermediate 2, and chemical reaction formula is as follows:
[0049]
[0050] The specific operation process is as follows: add 210g (1.71mol) of IIb 4-pyridinecarboxylic acid into a 2L reaction flask, then add 560mL of acetic acid and 210mL of 30% hydrogen peroxide, heat up to 90°C for 3 hours, then add 210mL of hydrogen peroxide, and continue the reaction for 3 hours. The reaction of the raw material 4-pyridinecarboxylic acid was detected by thin-layer chromatography. Slowly cool down to room temperature and add 210mL of acetone, then cool down to 0-5°C and stir for crystallization for 4h, filter, rinse the filter cake with a small amount of methyl tert-butyl ether, Air blowing dried to obtain 213 g of white solid with a yield of 90% and a purity of 99%.
[0051] Mass Spectrometry [M+H] + =140.2, nuclear magnetic detection 1H NMR (400MHz, DMSO) δ: 13.553 (br, 1H), 8.285-8.303 (dd, 2H), 7.816-7.834 (dd, 2H)....